Design of the First-in-Class, Highly Potent Irreversible Inhibitor Targeting the Menin-MLL Protein-Protein Interaction
Angew. Chem. Int. Ed., 2018
DOI: 10.1002/anie.201711828
Shaomeng Wang, Shilin Xu, Angelo Aguilar, Tianfeng Xu, Ke Zheng, Liyue Huang, Jeanne Stuckey, Krishnapriya Chinnaswamy, Denzil Bernard, Ester Fernández-Salas, Liu Liu, Mi Wang, Donna McEachern, Sally Przybranowski, Caroline Foster
We report the structure-based design of M-525 as the first-in-class, highly potent, irreversible small-molecule inhibitor of the menin-MLL interaction. M-525 targets cellular menin protein at sub-nanomolar concentrations and achieves low nanomolar potencies in cell growth inhibition and in suppression of MLL-regulated gene expression in MLL leukemia cells. M-525 demonstrates high cellular specificity over non-MLL leukemia cells and is >30-times more potent than its corresponding reversible inhibitors. Mass spectroscopic analysis and co-crystal structure of M-525 in complex with menin firmly establish its mode of action. A single administration of M-525 effectively suppresses MLL-regulated gene expression in tumor tissue. An efficient procedure was developed to synthesize M-525. Our study demonstrates that irreversible inhibition of menin may represent a promising therapeutic strategy for MLL leukemia.
Monday, January 1, 2018
Sunday, December 31, 2017
“Inverse Drug Discovery” Strategy To Identify Proteins That Are Targeted by Latent Electrophiles As Exemplified by Aryl Fluorosulfates
David E. Mortenson, Gabriel J. Brighty, Lars Plate, Grant Bare, Wentao Chen, Suhua Li, Hua Wang, Benjamin F. Cravatt, Stefano Forli , Evan T. Powers, K. Barry Sharpless Ian A. Wilson, and Jeffery W. Kelly
J. Am. Chem. Soc., Article ASAP
DOI: 10.1021/jacs.7b08366
Drug candidates are generally discovered using biochemical screens employing an isolated target protein or by utilizing cell-based phenotypic assays. Both noncovalent and covalent hits emerge from such endeavors. Herein, we exemplify an “Inverse Drug Discovery” strategy in which organic compounds of intermediate complexity harboring weak, but activatable, electrophiles are matched with the protein(s) they react with in cells or cell lysate. An alkyne substructure in each candidate small molecule enables affinity chromatography–mass spectrometry, which produces a list of proteins that each distinct compound reacts with. A notable feature of this approach is that it is agnostic with respect to the cellular proteins targeted. To illustrate this strategy, we employed aryl fluorosulfates, an underexplored class of sulfur(VI) halides, that are generally unreactive unless activated by protein binding. Reversible aryl fluorosulfate binding, correct juxtaposition of protein side chain functional groups, and transition-state stabilization of the S(VI) exchange reaction all seem to be critical for conjugate formation. The aryl fluorosulfates studied thus far exhibit chemoselective reactivity toward Lys and, particularly, Tyr side chains, and can be used to target nonenzymes (e.g., a hormone carrier or a small-molecule carrier protein) as well as enzymes. The “Inverse Drug Discovery” strategy should be particularly attractive as a means to explore latent electrophiles not typically used in medicinal chemistry efforts, until one reacts with a protein target of exceptional interest. Structure–activity data can then be used to enhance the selectivity of conjugate formation or the covalent probe can be used as a competitor to develop noncovalent drug candidates. Here we use the “Inverse Drug Discovery” platform to identify and validate covalent ligands for 11 different human proteins. In the case of one of these proteins, we have identified and validated a small-molecule probe for the first time.
J. Am. Chem. Soc., Article ASAP
DOI: 10.1021/jacs.7b08366
Drug candidates are generally discovered using biochemical screens employing an isolated target protein or by utilizing cell-based phenotypic assays. Both noncovalent and covalent hits emerge from such endeavors. Herein, we exemplify an “Inverse Drug Discovery” strategy in which organic compounds of intermediate complexity harboring weak, but activatable, electrophiles are matched with the protein(s) they react with in cells or cell lysate. An alkyne substructure in each candidate small molecule enables affinity chromatography–mass spectrometry, which produces a list of proteins that each distinct compound reacts with. A notable feature of this approach is that it is agnostic with respect to the cellular proteins targeted. To illustrate this strategy, we employed aryl fluorosulfates, an underexplored class of sulfur(VI) halides, that are generally unreactive unless activated by protein binding. Reversible aryl fluorosulfate binding, correct juxtaposition of protein side chain functional groups, and transition-state stabilization of the S(VI) exchange reaction all seem to be critical for conjugate formation. The aryl fluorosulfates studied thus far exhibit chemoselective reactivity toward Lys and, particularly, Tyr side chains, and can be used to target nonenzymes (e.g., a hormone carrier or a small-molecule carrier protein) as well as enzymes. The “Inverse Drug Discovery” strategy should be particularly attractive as a means to explore latent electrophiles not typically used in medicinal chemistry efforts, until one reacts with a protein target of exceptional interest. Structure–activity data can then be used to enhance the selectivity of conjugate formation or the covalent probe can be used as a competitor to develop noncovalent drug candidates. Here we use the “Inverse Drug Discovery” platform to identify and validate covalent ligands for 11 different human proteins. In the case of one of these proteins, we have identified and validated a small-molecule probe for the first time.

Saturday, December 16, 2017
Selectively targeting the kinome-conserved lysine of PI3Kδ as a general approach to covalent kinase inhibition
Samuel E Dalton, Lars Dittus, Daniel A. Thomas, Maire A. Convery, Joao Nunes, Jacob T. Bush, John P. Evans, Thilo Werner, Marcus Bantscheff, John A. Murphy, and Sebastien Campos
J. Am. Chem. Soc., 2017
DOI: 10.1021/jacs.7b08979
Selective covalent inhibition of kinases by targeting poorly conserved cysteines has proven highly fruitful to date in the development of chemical probes and approved drugs. However, this approach is limited to ~200 kinases possessing such a cysteine near the ATP-binding pocket. Herein, we report a novel approach to achieve selective, irreversible kinase inhibition, by targeting the conserved catalytic lysine residue. We have illustrated our approach by developing selective, covalent PI3Kδ inhibitors that exhibit nanomolar potency in cellular assays, and a duration of action >48 h in CD4+ T cells. Despite conservation of the lysine residue throughout the kinome, the lead compound shows high levels of selectivity over a selection of lipid and protein kinases in biochemical assays, as well as covalent binding to very few off-target proteins in live-cell proteomic studies. We anticipate this approach could offer an alternative general strategy, to targeting non-conserved cysteines, for the development of selective covalent kinase inhibitors.
J. Am. Chem. Soc., 2017
DOI: 10.1021/jacs.7b08979
Selective covalent inhibition of kinases by targeting poorly conserved cysteines has proven highly fruitful to date in the development of chemical probes and approved drugs. However, this approach is limited to ~200 kinases possessing such a cysteine near the ATP-binding pocket. Herein, we report a novel approach to achieve selective, irreversible kinase inhibition, by targeting the conserved catalytic lysine residue. We have illustrated our approach by developing selective, covalent PI3Kδ inhibitors that exhibit nanomolar potency in cellular assays, and a duration of action >48 h in CD4+ T cells. Despite conservation of the lysine residue throughout the kinome, the lead compound shows high levels of selectivity over a selection of lipid and protein kinases in biochemical assays, as well as covalent binding to very few off-target proteins in live-cell proteomic studies. We anticipate this approach could offer an alternative general strategy, to targeting non-conserved cysteines, for the development of selective covalent kinase inhibitors.
Wednesday, December 6, 2017
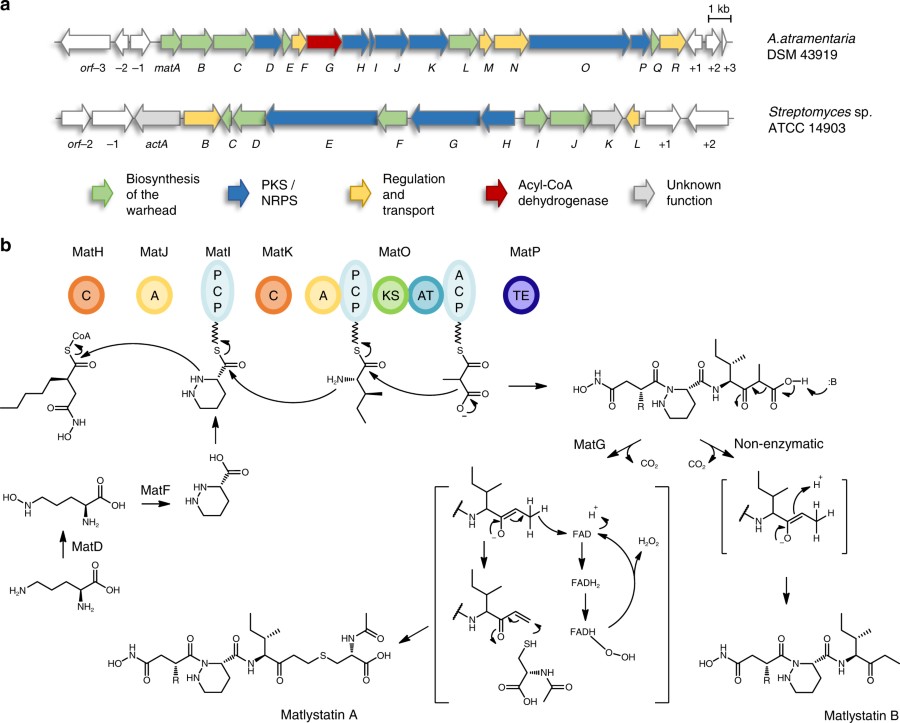
Warhead biosynthesis and the origin of structural diversity in hydroxamate metalloproteinase inhibitors
Franziska Leipoldt, Javier Santos-Aberturas, Dennis P. Stegmann, Felix Wolf, Andreas Kulik, Rodney Lacret, Désirée Popadić, Daniela Keinhörster, Norbert Kirchner, Paulina Bekiesch, Harald Gross, Andrew W. Truman & Leonard Kaysser
doi: 10.1038/s41467-017-01975-6
Metalloproteinase inhibitors often feature hydroxamate moieties to facilitate the chelation of metal ions in the catalytic center of target enzymes. Actinonin and matlystatins are potent metalloproteinase inhibitors that comprise rare N-hydroxy-2-pentyl-succinamic acid warheads. Here we report the identification and characterization of their biosynthetic pathways. By gene cluster comparison and a combination of precursor feeding studies, heterologous pathway expression and gene deletion experiments we are able to show that the N-hydroxy-alkyl-succinamic acid warhead is generated by an unprecedented variation of the ethylmalonyl-CoA pathway. Moreover, we present evidence that the remarkable structural diversity of matlystatin congeners originates from the activity of a decarboxylase-dehydrogenase enzyme with high similarity to enzymes that form epoxyketones. We further exploit this mechanism to direct the biosynthesis of non-natural matlystatin derivatives. Our work paves the way for follow-up studies on these fascinating pathways and allows the identification of new protease inhibitors by genome mining.
Tuesday, December 5, 2017
Chemoselective Installation of Amine Bonds on Proteins through Aza-Michael Ligation
Allyson M. Freedy, Maria J. Matos, Omar Boutureira, Francisco Corzana, Ana Guerreiro, Padma Akkapeddi, Víctor J. Somovilla, Tiago Rodrigues, Karl Nicholls, Bangwen Xie, Gonzalo Jiménez-Osés, Kevin M. Brindle, André A. Neves, & Gonçalo J. L. Bernardes
J. Am. Chem. Soc., 2017 ASAP
DOI: 10.1021/jacs.7b10702
Chemical modification of proteins is essential for a variety of important diagnostic and therapeutic applications. Many strategies developed to date lack chemo- and regioselectivity as well as result in non-native linkages that may suffer from instability in vivo and adversely affect the protein’s structure and function. We describe here the reaction of N-nucleophiles with the amino acid dehydroalanine (Dha) in a protein context. When Dha is chemically installed in proteins, the addition of a wide-range N-nucleophiles enables the rapid formation of amine linkages (secondary and tertiary) in a chemoselective manner under mild, biocompatible conditions. These new linkages are stable at a wide range of pH values (pH 2.8 to 12.8), under reducing conditions (biological thiols such as glutathione) and in human plasma. This method is demonstrated for three proteins and is shown to be fully compatible with disulfide bridges, as evidenced by the selective modification of recombinant albumin that displays 17 structurally relevant disulfides. The practicability and utility of our approach is further demonstrated by the construction of a chemically modified C2A domain of Synaptotagmin-I protein that retains its ability to preferentially bind to apoptotic cells at a level comparable to the native protein. Importantly, the method was useful for building a homogeneous antibody-drug conjugate with a precise drug-to-antibody ratio of 2. The kinase inhibitor crizotinib was directly conjugated to Dha through its piperidine motif, and its antibody-mediated intracellular delivery results in 10-fold improvement of its cancer cell-killing efficacy. The simplicity and exquisite site-selectivity of the aza-Michael ligation described herein allows the construction of stable secondary and tertiary amine-linked protein conjugates without affecting the structure and function of biologically relevant proteins.

Friday, November 17, 2017
Covalent Ligand Discovery against Druggable Hotspots Targeted by Anti-cancer Natural Products
Elizabeth A. Grossman, Carl C. Ward, Jessica N. Spradlin, Leslie A. Bateman, Tucker R. Huffman, David K. Miyamoto, Jordan I. Kleinman, Daniel K. Nomura
Cell Chemical Biology, 2017
doi: 10.1016/j.chembiol.2017.08.013
Many natural products that show therapeutic activities are often difficult to synthesize or isolate and have unknown targets, hindering their development as drugs. Identifying druggable hotspots targeted by covalently acting anti-cancer natural products can enable pharmacological interrogation of these sites with more synthetically tractable compounds. Here, we used chemoproteomic platforms to discover that the anti-cancer natural product withaferin A targets C377 on the regulatory subunit PPP2R1A of the tumor-suppressor protein phosphatase 2A (PP2A) complex leading to activation of PP2A activity, inactivation of AKT, and impaired breast cancer cell proliferation. We developed a more synthetically tractable cysteine-reactive covalent ligand, JNS 1-40, that selectively targets C377 of PPP2R1A to impair breast cancer signaling, proliferation, and in vivo tumor growth. Our study highlights the utility of using chemoproteomics to map druggable hotspots targeted by complex natural products and subsequently interrogating these sites with more synthetically tractable covalent ligands for cancer therapy.

Cell Chemical Biology, 2017
doi: 10.1016/j.chembiol.2017.08.013
Many natural products that show therapeutic activities are often difficult to synthesize or isolate and have unknown targets, hindering their development as drugs. Identifying druggable hotspots targeted by covalently acting anti-cancer natural products can enable pharmacological interrogation of these sites with more synthetically tractable compounds. Here, we used chemoproteomic platforms to discover that the anti-cancer natural product withaferin A targets C377 on the regulatory subunit PPP2R1A of the tumor-suppressor protein phosphatase 2A (PP2A) complex leading to activation of PP2A activity, inactivation of AKT, and impaired breast cancer cell proliferation. We developed a more synthetically tractable cysteine-reactive covalent ligand, JNS 1-40, that selectively targets C377 of PPP2R1A to impair breast cancer signaling, proliferation, and in vivo tumor growth. Our study highlights the utility of using chemoproteomics to map druggable hotspots targeted by complex natural products and subsequently interrogating these sites with more synthetically tractable covalent ligands for cancer therapy.
Tuesday, November 14, 2017
Can Relative Binding Free Energy Predict Selectivity of Reversible Covalent Inhibitors?
Payal Chatterjee, Wesley M. Botello-Smith, Han Zhang, Li Qian, Abdelaziz Alsamarah, David Kent, Jerome J. Lacroix, Michel Baudry, and Yun Luo
J. Am. Chem. Soc., 2017
doi: 10.1021/jacs.7b08938
Reversible covalent inhibitors have many clinical advantages over noncovalent or covalent drugs. However, apart from selecting a warhead, substantial efforts in design and synthesis are needed to optimize noncovalent interactions to improve target-selective binding. Computational prediction of binding affinity for reversible covalent inhibitors presents a unique challenge since the binding process consists of multiple steps, which are not necessarily independent of each other. In this study, we lay out the relation between relative binding free energy and the overall reversible covalent binding affinity using a two-state binding model. To prove the concept, we employed free energy perturbation (FEP) coupled with λ-exchange molecular dynamics method to calculate the binding free energy of a series of α-ketoamide analogs relative to a common warhead scaffold, in both noncovalent and covalent bond states, and for two highly homologous proteases, calpain-1 and calpain-2. We conclude that covalent binding affinity alone, in general, can be used to predict reversible covalent binding selectivity. However, exceptions may exist. Therefore, we also discuss the conditions under which the noncovalent binding step is no longer negligible and propose a novel approach that combines the relative FEP calculations with a single QM/MM calculation of warhead to predict the binding affinity and binding kinetics for a large number of reversible covalent inhibitors. Our FEP calculations also revealed that covalent and noncovalent states of an inhibitor do not necessarily exhibit the same selectivity. Thus, investigating both binding states, as well as the kinetics will provide extremely useful information for optimizing reversible covalent inhibitors.
J. Am. Chem. Soc., 2017
doi: 10.1021/jacs.7b08938
Reversible covalent inhibitors have many clinical advantages over noncovalent or covalent drugs. However, apart from selecting a warhead, substantial efforts in design and synthesis are needed to optimize noncovalent interactions to improve target-selective binding. Computational prediction of binding affinity for reversible covalent inhibitors presents a unique challenge since the binding process consists of multiple steps, which are not necessarily independent of each other. In this study, we lay out the relation between relative binding free energy and the overall reversible covalent binding affinity using a two-state binding model. To prove the concept, we employed free energy perturbation (FEP) coupled with λ-exchange molecular dynamics method to calculate the binding free energy of a series of α-ketoamide analogs relative to a common warhead scaffold, in both noncovalent and covalent bond states, and for two highly homologous proteases, calpain-1 and calpain-2. We conclude that covalent binding affinity alone, in general, can be used to predict reversible covalent binding selectivity. However, exceptions may exist. Therefore, we also discuss the conditions under which the noncovalent binding step is no longer negligible and propose a novel approach that combines the relative FEP calculations with a single QM/MM calculation of warhead to predict the binding affinity and binding kinetics for a large number of reversible covalent inhibitors. Our FEP calculations also revealed that covalent and noncovalent states of an inhibitor do not necessarily exhibit the same selectivity. Thus, investigating both binding states, as well as the kinetics will provide extremely useful information for optimizing reversible covalent inhibitors.
Subscribe to:
Posts (Atom)